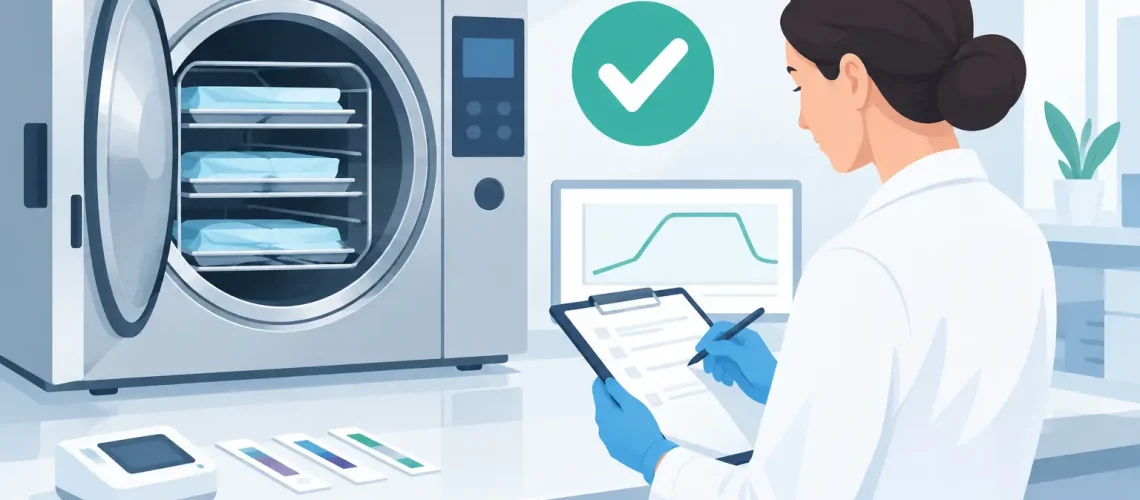
A sterilizer can reach its programmed temperature and still fail to sterilize the most difficult item in the load. Dense packs, trapped air, poor steam penetration, overloaded trolleys, cold spots, and incorrect packaging can all change the result. Knowing how to validate sterilization cycles means proving that a specific machine, cycle, load configuration, and operating procedure consistently achieve the required microbial kill.
For hospitals, laboratories, food operations, laundries, and industrial facilities, validation is not a one-time temperature check. It is an evidence-based process that connects equipment capability with the real conditions of daily operation. The objective is straightforward: demonstrate that every critical point in the defined load receives the required sterilizing exposure, then maintain that performance through monitoring, maintenance, and documented control.
Start With the Process You Need to Prove
Validation begins before instruments are placed inside a chamber. Define what is being sterilized, the intended sterility assurance requirement, the sterilization method, the load type, and the acceptance criteria. A cycle designed for unwrapped metal tools may not be suitable for porous textiles, liquid containers, hollow devices, or tightly packed production components.
The sterilization method determines the variables that matter most. Steam sterilization depends on saturated steam, effective air removal, temperature, pressure, and exposure time. Dry-heat sterilization depends heavily on temperature uniformity, air circulation, loading density, and exposure duration. Chemical, low-temperature, and other specialized processes require their own validated parameters and compatible indicators.
The equipment manufacturer’s instructions, your internal quality procedures, customer specifications, and applicable regulations should all inform the protocol. Do not copy a cycle setting from a different facility and assume it will perform the same way. Chamber size, steam supply quality, utility stability, rack design, packaging, and product geometry all affect process results.
How to Validate Sterilization Cycles Step by Step
Confirm the equipment is qualified
Before validating a cycle, confirm the sterilizer and its supporting utilities are operating as designed. This usually includes installation qualification, operational qualification, and performance qualification.
Installation qualification verifies that the machine has been installed correctly. Check electrical supply, steam or water connections where applicable, drainage, ventilation, safety interlocks, chamber seals, sensors, control panels, and calibration status. For custom-built equipment, this stage should also confirm that the agreed chamber configuration, trolley arrangement, controls, and safety features match the approved design.
Operational qualification verifies that the equipment can run across its intended operating range. Test alarms, door interlocks, cycle controls, recording functions, emergency stops, and programmed setpoints. If a sensor fails, a door does not seal, or a controller records inaccurate data, a successful biological test alone does not establish a dependable process.
Performance qualification demonstrates that the sterilizer performs under real or simulated production conditions. This is where the defined load, worst-case locations, and biological challenge become central.
Identify the worst-case load and cold spots
The hardest load to sterilize should be the focus of validation. In many operations, that means the heaviest load, the densest package arrangement, the longest trolley, the largest container, or the product location furthest from effective heat or steam circulation.
Map the chamber with calibrated temperature sensors or data loggers. Place them at locations likely to heat slowly or receive limited steam access, such as corners, lower racks, the center of dense loads, and areas near the door. For larger trolley sterilizers or custom industrial chambers, mapping should reflect the actual trolley dimensions and loading pattern rather than an empty-chamber test alone.
Empty-chamber mapping is useful for understanding machine behavior, but it cannot substitute for loaded-cycle validation. Product mass and packaging can create thermal lag and obstruct circulation. A cycle that passes when empty may be inadequate once production loads are introduced.
Use three forms of process evidence
A validated cycle normally relies on physical, chemical, and biological evidence. Each has a different role.
Physical monitoring comes from the sterilizer’s validated sensors, independent data loggers, chart recorders, or electronic batch records. These records show whether time, temperature, pressure, and other programmed variables were achieved. Sensors must be calibrated against traceable references at a defined frequency. A display reading is only useful if the underlying measurement system is accurate.
Chemical indicators respond to one or more sterilization conditions. They are valuable for distinguishing processed from unprocessed items and for identifying potential exposure problems at the package or load level. However, they do not prove sterility by themselves. Use them according to the indicator manufacturer’s instructions and your facility procedure.
Biological indicators provide the most direct challenge because they contain highly resistant test microorganisms appropriate to the sterilization method. Place them in the locations least likely to receive the required exposure, including representative internal areas of challenging packs or process challenge devices. After the cycle, incubate and interpret them according to the biological indicator manufacturer’s instructions. A negative result supports the conclusion that the defined process achieved the required lethality.
For pre-vacuum steam sterilizers, air-removal testing may also be required. Ineffective air removal can prevent saturated steam from contacting surfaces even when chamber temperature appears acceptable. The right test method depends on the equipment design and the applicable standard or facility requirement.
Run repeat cycles, not a single successful test
One acceptable cycle may show that the machine can work. Repeated successful cycles show that the process is reproducible. Many validation protocols use multiple consecutive runs with the same approved worst-case load. The precise number of runs and acceptance criteria should be established by your quality system, risk assessment, and governing requirements.
During each run, record the load description, cycle recipe, sensor positions, physical data, chemical indicator results, biological indicator locations and outcomes, operator, date, utility conditions, and any deviations. Repetition is particularly important after major repairs, control-system changes, chamber modifications, relocation, or a new packaging and loading configuration.
Set Clear Acceptance Criteria Before Testing
Do not interpret results after the fact based on what looks reasonable. Write acceptance criteria before the validation starts. These criteria should state the required exposure conditions, allowable temperature variation, acceptable pressure behavior where relevant, biological indicator result, chemical indicator response, and documentation requirements.
For example, a steam process may require every mapped point to remain within a defined temperature range for the full exposure period, while all biological indicators in the challenge locations show no growth. A dry-heat process may place greater emphasis on temperature distribution, heat penetration, and exposure duration at the slowest-heating point.
Acceptance criteria must align with the validated method and intended application. More temperature is not automatically better. Excessive heat can damage products, packaging, seals, electronics, lubricants, or sensitive components. The right cycle is the one that achieves sterilization reliably without compromising product quality or safety.
Control the Routine Process After Validation
Validation establishes a baseline. Routine monitoring keeps the process under control. Every load should have a traceable record that confirms the correct cycle was selected and the required parameters were met. Depending on the process, this may include digital batch records, physical chart records, chemical indicators, load labels, and operator signoff.
Preventive maintenance is part of sterilization assurance. Worn door gaskets, blocked drains, damaged racks, unstable steam quality, failing valves, and drifting sensors can gradually reduce performance. A scheduled maintenance plan should cover mechanical inspection, cleaning, calibration, utility checks, alarm tests, and replacement of wear components.
Facility teams should also control loading practices. Define maximum load sizes, approved packaging materials, trolley orientation, spacing requirements, and placement rules. A validated cycle can be invalidated by an operator placing items too tightly together or introducing a product that was never assessed during qualification.
For custom sterilization machinery, this is where engineering support adds practical value. REK Engineering Machinery can design equipment around a facility’s process needs, but the operating team must still validate the final machine within its actual environment, utilities, load types, and quality procedures.
Know When Revalidation Is Required
Revalidation is not reserved for complete equipment replacement. It should be considered whenever a change could affect heat transfer, steam penetration, time-temperature exposure, sensor accuracy, or load configuration. Common triggers include controller upgrades, chamber repairs, relocation, utility changes, altered packaging, new product types, revised loading patterns, and repeated monitoring failures.
A risk-based change-control process helps determine whether a full revalidation or a targeted verification is appropriate. Replacing a noncritical exterior panel may need only maintenance documentation. Replacing a temperature sensor, modifying a trolley, or changing the cycle recipe demands far more scrutiny.
If a cycle fails, quarantine the affected load where required, investigate the cause, correct the issue, and document the decision. Do not simply rerun the cycle without understanding whether the problem was an indicator error, operator mistake, utility interruption, equipment fault, or a genuine process failure.
A well-validated sterilization process gives operators a clear standard to follow and gives management evidence that hygiene controls are working as intended. Treat each validation as a working engineering document: specific enough for the people running the equipment, disciplined enough for audits, and flexible enough to guide safe action when the process changes.